Feeding the Brain When Energy Runs Low
Dr. Nayiar Shahid - 20 April 2026
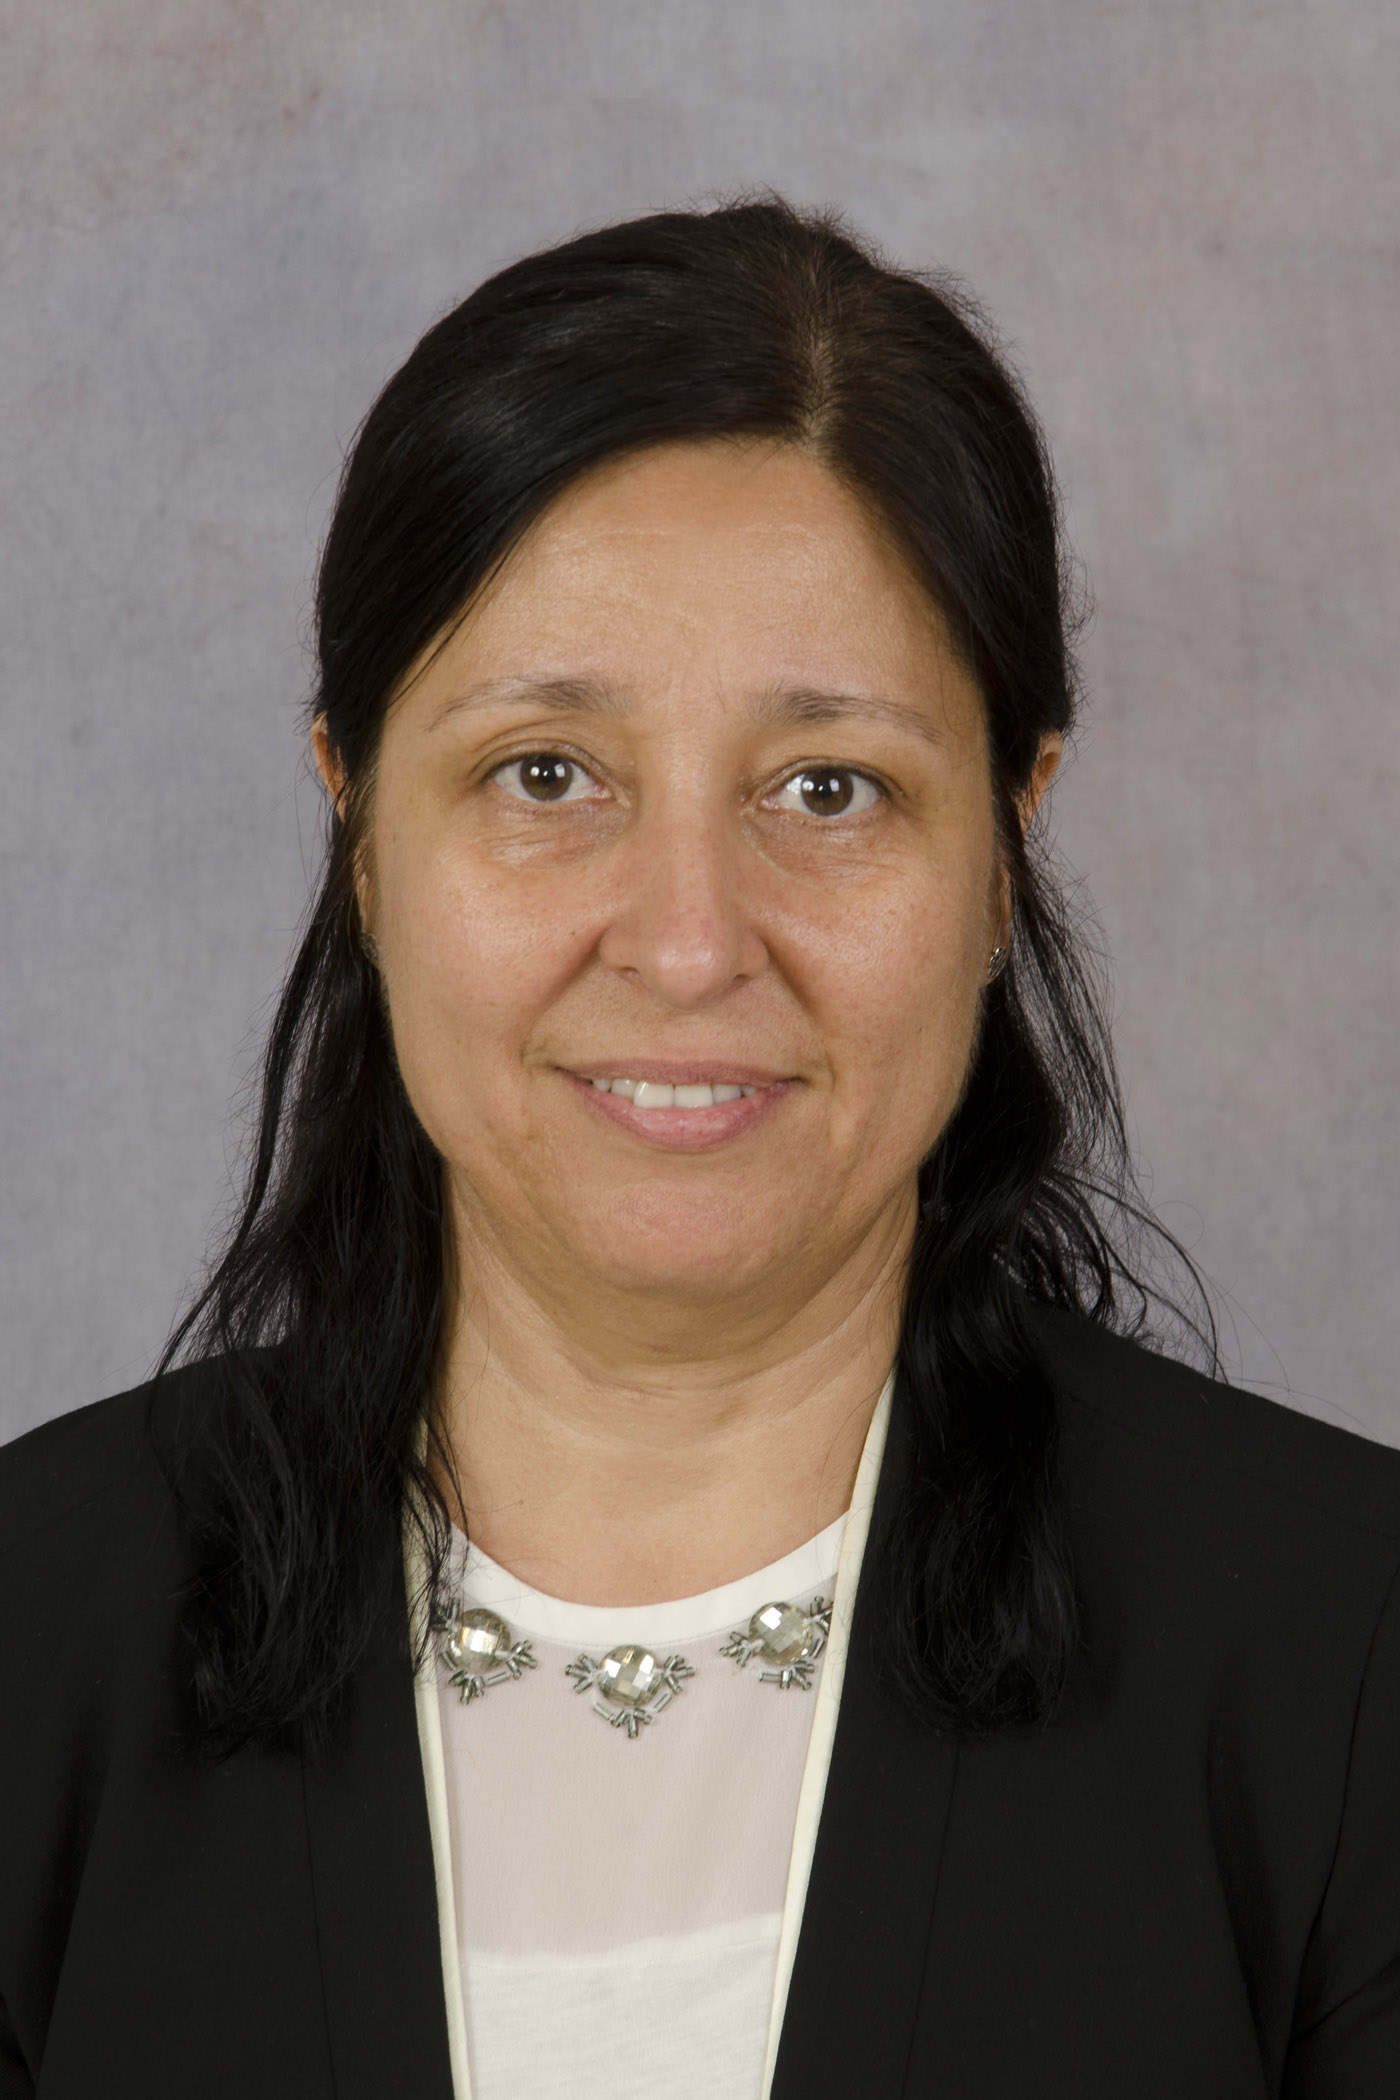
Dr. Saadet Mercimek-Andrews
Some children are born with a rare condition that leads to seizures early in life. This disorder is called pyridoxine-dependent epilepsy (PDE) and it is caused by changes in the ALDH7A1 gene, which codes for the enzymatic protein, aldehyde dehydrogenase 7 family member A1. The enzyme is responsible for breaking down a toxic chemical in the body. When this enzyme does not function properly, the toxin builds up in the body and brain leading to numerous cellular and system-wide dysfunctions. Thankfully, the condition can be managed with the addition of vitamin B6, arginine and tryptophan in the diet and decreasing protein intake.
Recent work in this area has demonstrated that the dysfunction may be energy-based. Specifically, one step in energy production known as the tricarboxylic acid or TCA cycle - you might remember it as the Krebs cycle from school - is disrupted, reducing the amount of energy produced in the body.
This has led Dr. Saadet Andrews to explore alternate options to better treat these children. The goal is to use a molecule known to help the TCA cycle. It’s called triheptanoin and it was the treatment used in a n-of-1 clinical trial now published in Therapeutic Advances in Rare Disease.
Triheptanoin is a special type of fat, sometimes described as a medical oil. Unlike regular fats, it can be quickly converted into molecules that refill the TCA cycle. This process, called anaplerosis, helps restore the cell’s ability to produce energy.
The study followed a 4-year-old boy whose development had remained severely limited despite standard treatments of PDE. After adding triheptanoin to his therapy, Dr. Andrews' research team observed a meaningful change.
Over time, his cognitive development improved, rising from about 16% to 63% of expected levels for his age. This kind of improvement is unusual in PDE, where progress is often slow even after seizures are controlled.
Importantly, the treatment was also well tolerated, with no major safety concerns reported during the study period. The child did have nausea and retching for up to three weeks after dose increases, but this improved and allowed the dose to be increased gradually.
While this is only a single case, the results open the door to a new way of thinking about treatment. Instead of focusing only on stopping seizures or reducing toxins, therapies could also aim to support energy needs directly. This falls in line with previous work that has demonstrated the importance of energy efficiency in other children. While there is no doubt more research is needed to confirm these results in larger groups of patients, this study provides early evidence that targeting energy metabolism could make a meaningful difference.
In rare diseases like PDE, even one successful case can point researchers toward entirely new strategies. Sometimes, helping the brain isn’t just about stopping what’s going wrong.. it’s about giving it the fuel it needs to move forward.